Devices used to change rarely. Now updates ship all year, and every one is a design change. The paperwork fits the old pace, and the releases have stopped waiting for it.
Quality and compliance for medical device companiesFrom ISO 13485
From ISO 13485
to market clearance
Design controls, the ISO 14971 risk file and the technical documentation, alongside the procedures that govern them.

Medical device companies on Qualio, from Class II 510(k) to Class III, IVD and SaMD
EVERY RELEASE IS A DESIGN CHANGE
Your products are changing. Compliance needs to keep up.
The risk file trails the release
The hazard analysis covers the last version, not this one.
Training runs on the old revision
The procedure changed. The training records did not.
The technical file falls behind
Rebuilt by hand before every submission.
THE PAPERWORK IS THE AUTOMATED PART
Qualio compiles the technical file and drafts the reviews
Qualio reads the design controls as your engineers change them in Jira, drafts the review and the change control, and routes both to the approvers your procedure names. Once signed, the change goes into the technical documentation, compiled for 510(k), MDR and your market registrations, and Qualio reassesses the records it touched against ISO 13485 and the QMSR, so the documentation is current when the release ships.
The Results
6 months
To ISO 13485 certification.
Cardiaccs
510(k)
Cleared on a Qualio quality system.
EvoEndo42 → 0
Audit findings closed in 90 days.
CergenX
How it works
Your engineers stay in their tools. Qualio reads the work as it happens.
No plugin, no new Jira structure, no migration needed.
Quality Management
Qualio holds the procedures, training, complaints, CAPAs, change controls and supplier records the whole company works in, so every audit finds the approved version of each.
Regulated Product Lifecycle
Qualio links requirements, tests and code to the hazards and design inputs as the work changes, so the technical documentation compiles from design controls your approvers signed, for each market you register in.
Compliance Frameworks
Qualio maps ISO 13485, the QMSR, ISO 14971 and MDR once and assesses every record clause by clause as it changes, so the audit pack exports with the evidence behind each clause.
Copilot
Drafts the change control, the CAPA write-up, the document review and the training quiz from your own records. Every output cites its source and waits for your approval.
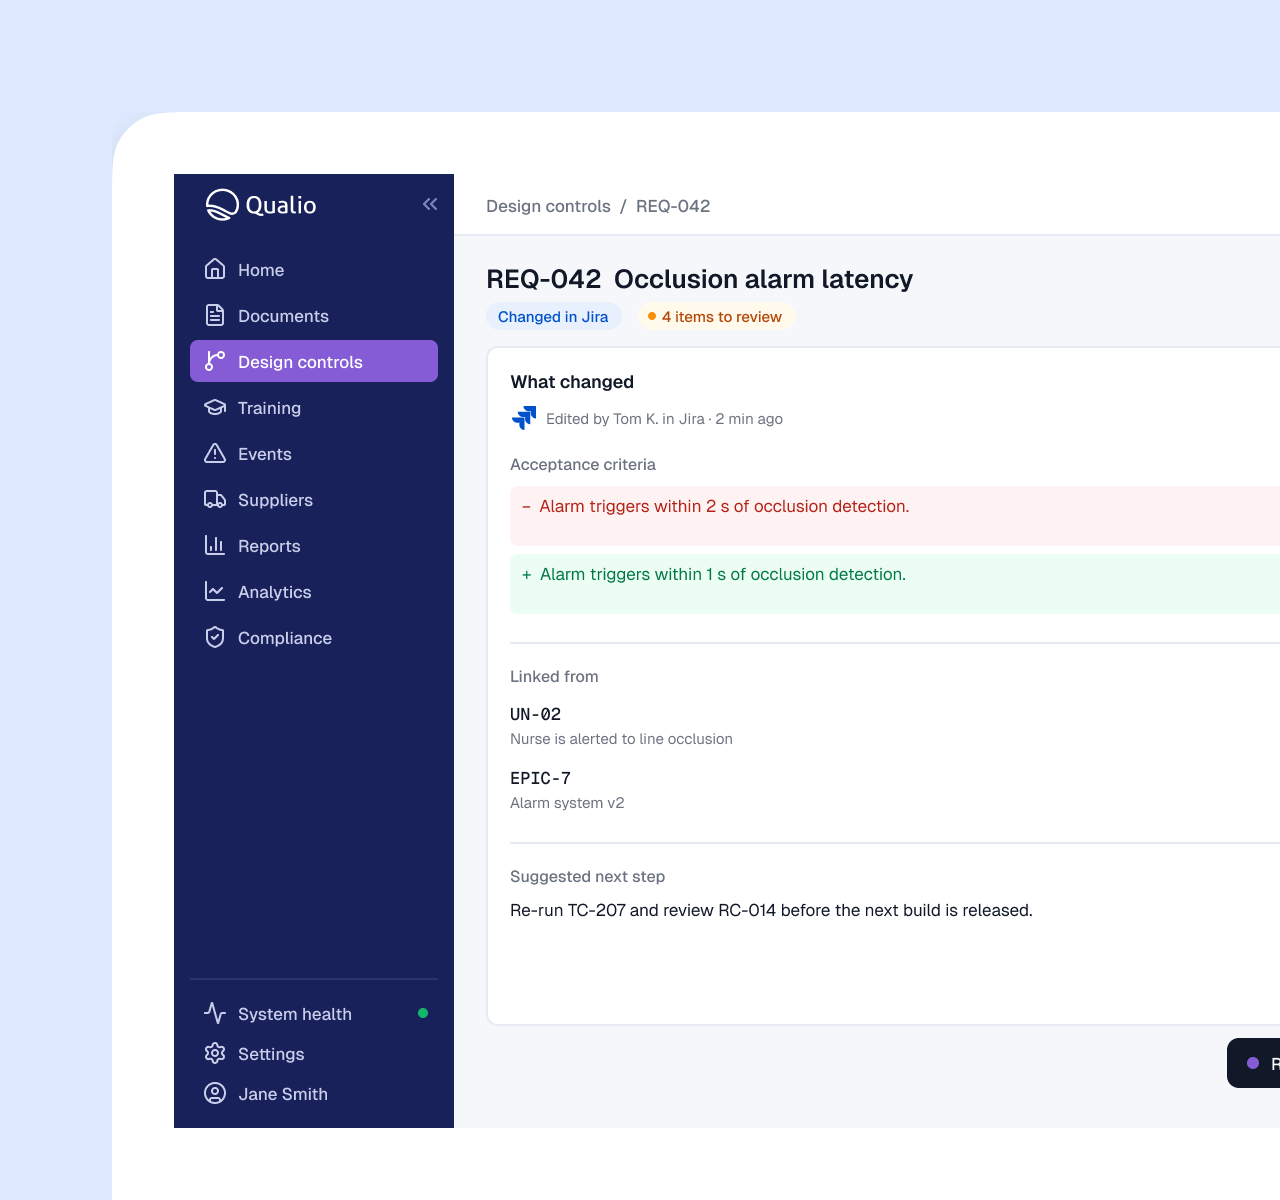
CHANGE CONTROL UNDER ISO 13485, THE QMSR AND IEC 62304
Every change shows what it touches before anyone approves it
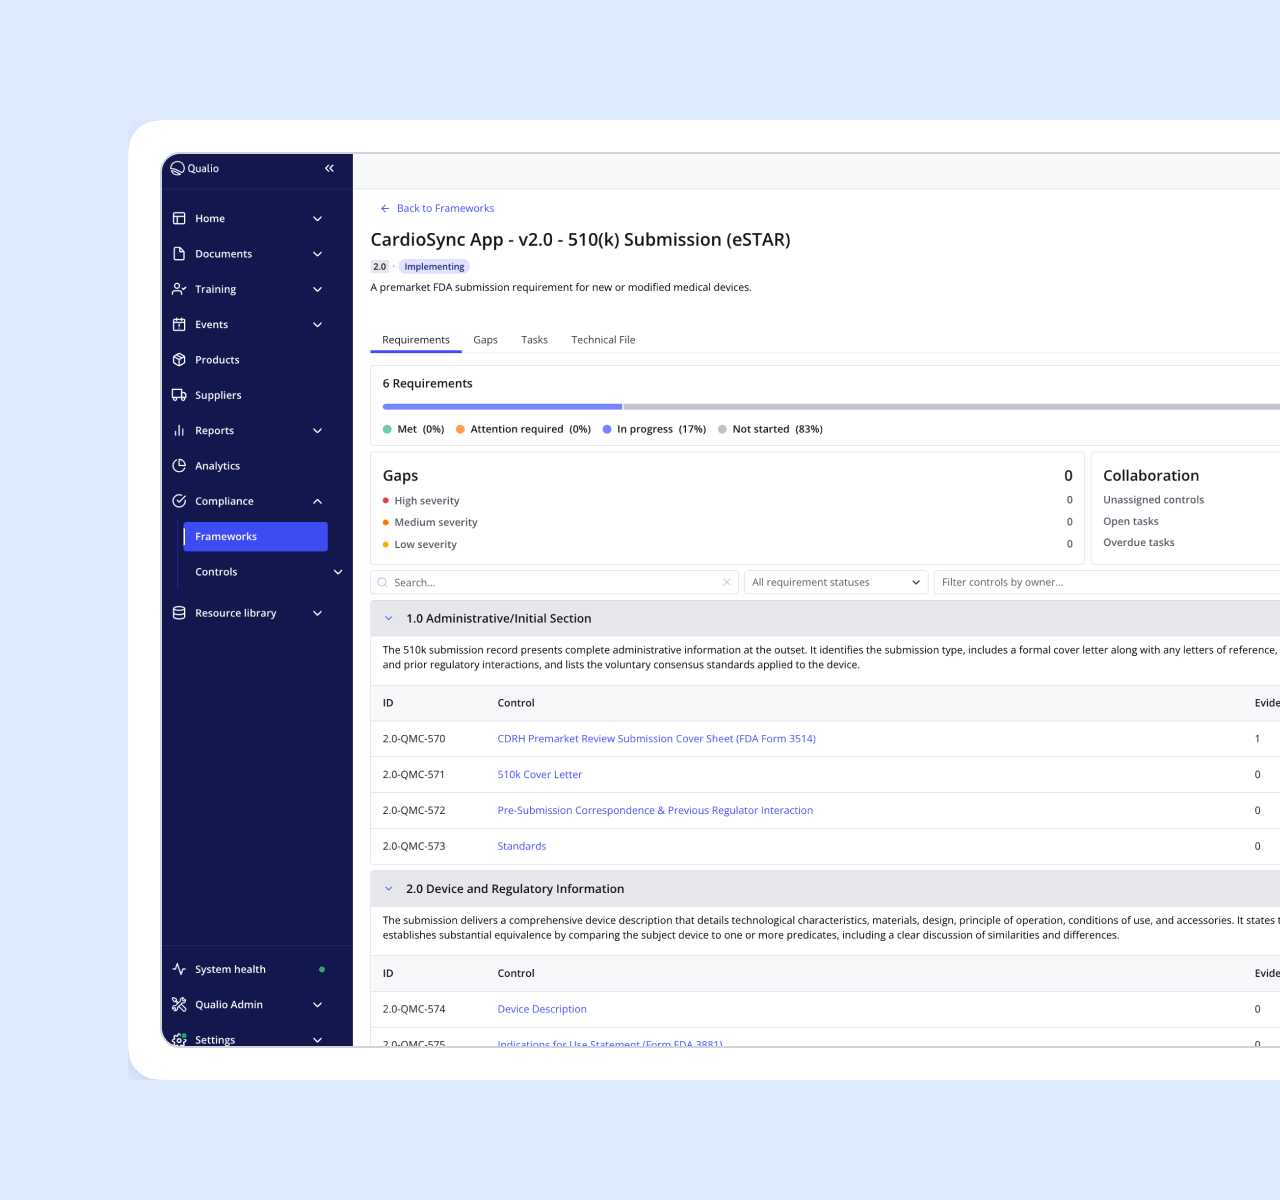
510(k) AND CE MARKING
The reviewer gets the whole technical file, with the approval behind every document
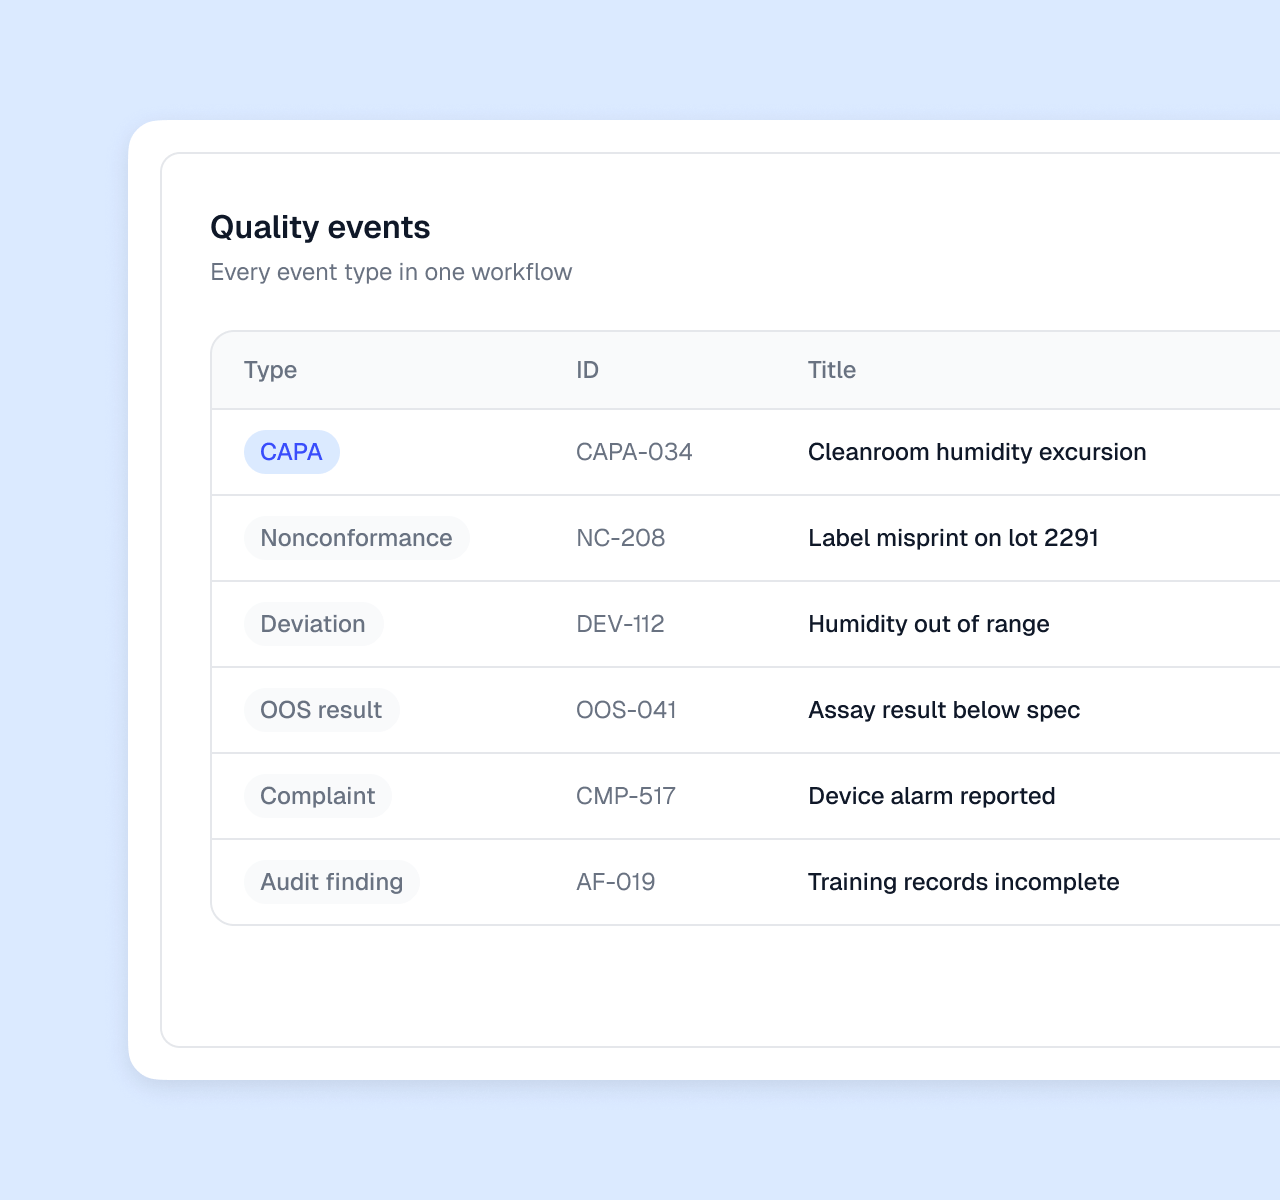
COMPLAINTS AND VIGILANCE
Qualio keeps the reportability decision with the complaint, dated and signed
What device companies say
“Just building a compendium of everything you need to show the notified bodies can be tricky if you're doing it manually. We wanted to start on the right footing.”
Daniel Aragao
CTO, InVivo Bionics
“Qualio's Design Controls feature has been an important element of our product development and regulatory approval processes. We provide an AI-based medical device that screens mammograms and detects anomalies. Quality and speed are critical because we know delays on our end impact real peoples' lives.”
Kevin Harris
President & Co-Founder, CureMetrix
“We now have a consolidated device file which reflects the development of our product. 18 months ago, we didn't have anything like that.”
Phil Cooley
Quality Manager, Bond Digital Health