Regulated Product LifecycleYour technical file
Your technical file
is your release
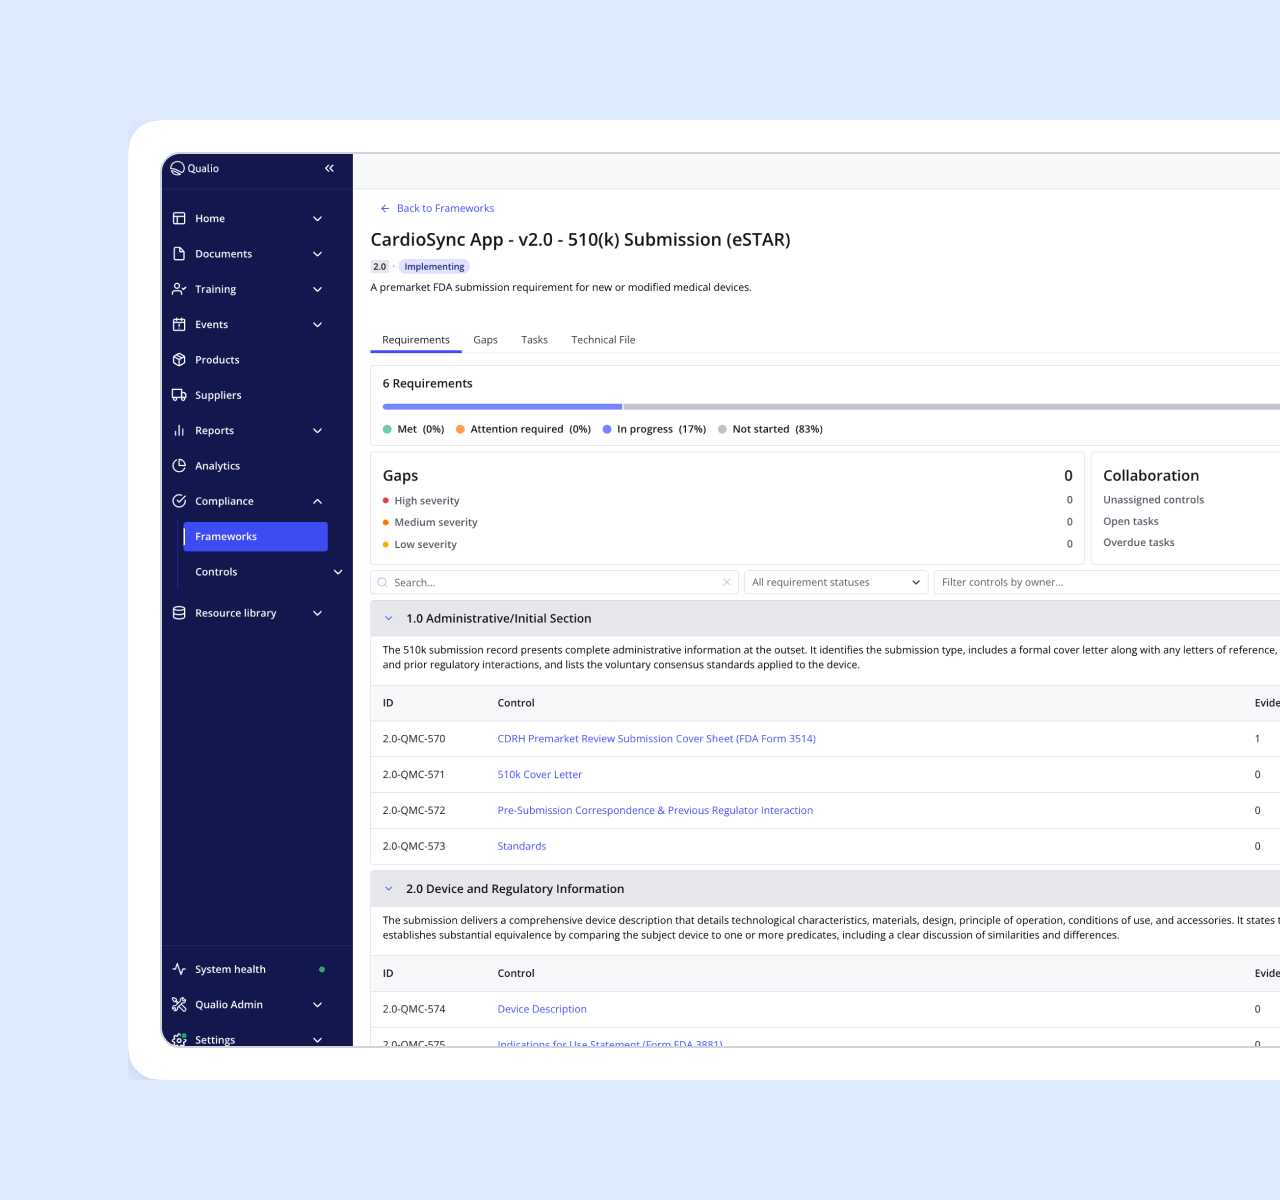
Build the 510(k), the EU MDR technical documentation and each market registration from the documents your approvers signed. One file per product version, frozen the day you submit.
700+ life sciences companies
4.7/5 on
Your technical documentation compiles from approved documents
What you submitted stays on file while the product moves on
Version 2.0 starts from the 1.0 baseline
Submit in multiple markets from the same documents
Platform connections
Design controls
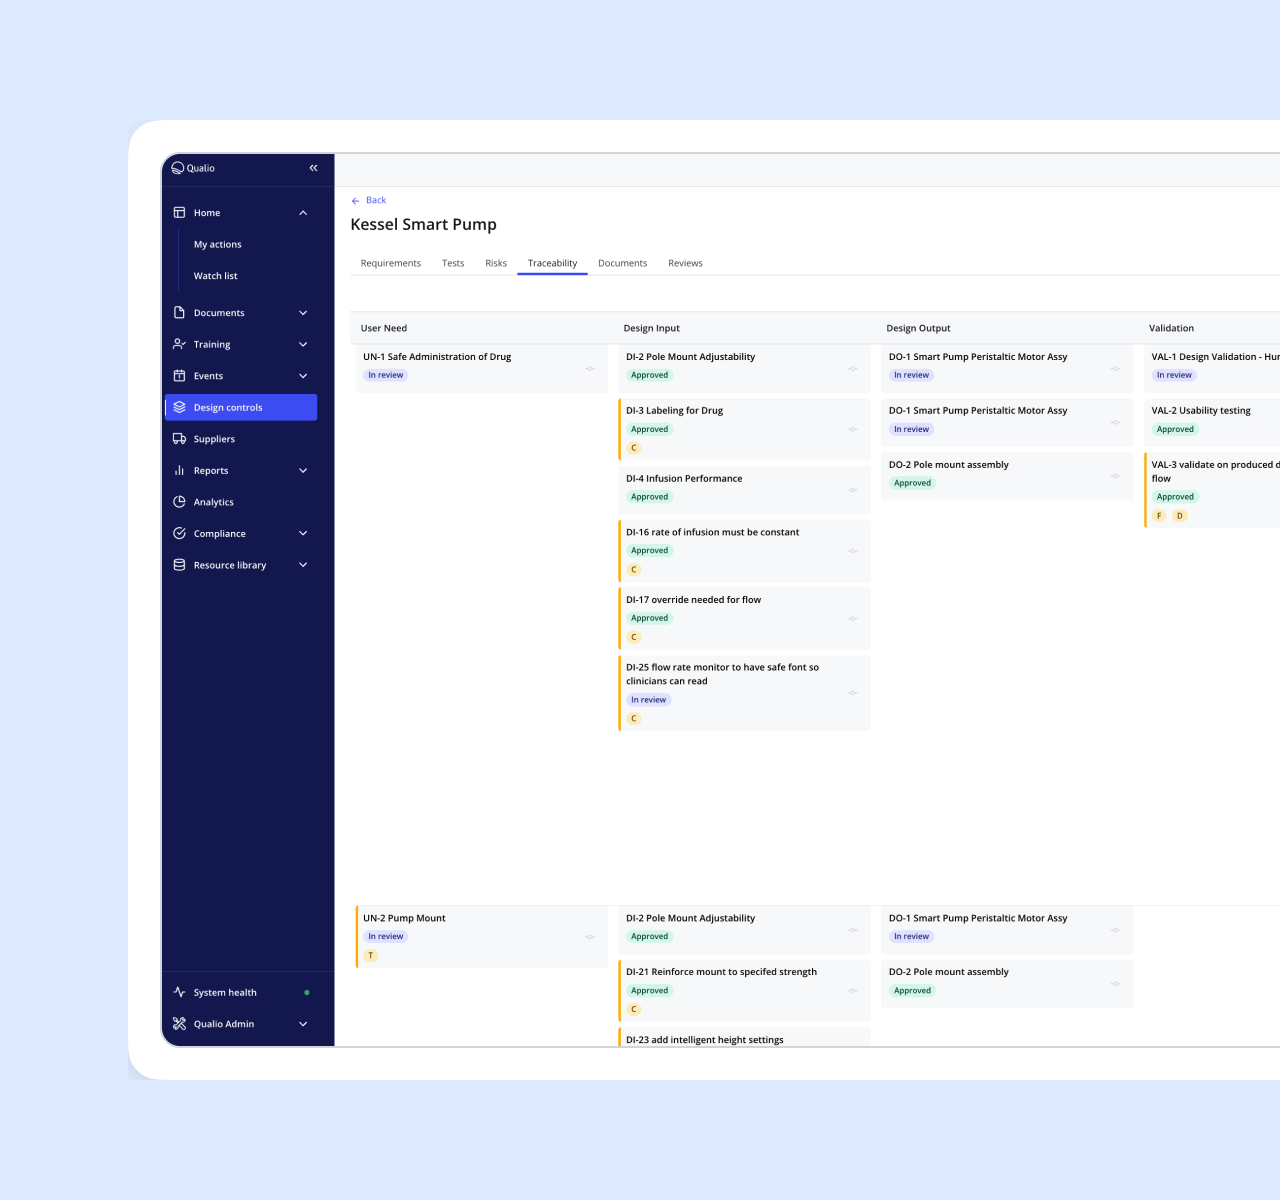
The package an approved design review generates, trace matrix and risk file included, is placed in the product version as a document, so it sits in the file at the version the approvers signed. See Design controls.
Compliance Frameworks
The 510(k) and EU MDR frameworks come from the same library as ISO 13485 and the FDA QMSR, so the file the reviewer reads and the system the auditor inspects are built the same way. See Compliance Frameworks.
Document control
Every document in the file is a controlled document with its own review, approval and audit trail. See Document control.
Frequently Asked Questions
What is a technical file, and is it the same as a DHF?
The technical file is the set of documents showing a device meets its requirements: design, risk management file, verification and validation, labelling and post-market surveillance. EU MDR Annex II and III set its contents. The FDA’s design history file was the design part of it. The QMSR now points to ISO 13485, which calls it the medical device file.
Is the export in eSTAR order or Annex II order?
It follows the framework you applied. Under the 510(k) framework the file is laid out in the eSTAR’s sections. Under the EU MDR framework it follows Annex II and Annex III. The download is one zip, index PDF first, or file by file. You still complete the FDA’s eSTAR form itself.
What does the notified body get, and can it change afterwards?
A snapshot: the file of one product version at the moment you took it, an index PDF listing every section and the document under it, plus one PDF of each document as it stood. It is read-only once generated. The documents inside carry their own approvals. When the file moves on, you take another.
What happens to the file when we release 2.0?
You create 2.0 with 1.0 as its baseline. Documents that run for the product’s life copy across, and the new file marks what changed against the baseline. Documents written for 1.0 stay on 1.0 and its snapshots, so what was valid for 1.0 is still there when the notified body or the FDA asks for it.
Do we need one framework per market?
No. The 510(k) and EU MDR frameworks share controls, so a document placed under a shared control appears in both. For a market with no framework in the library, build one in the framework builder from the regulator’s requirements and apply it to the version you register there. Not applicable is allowed on any section.
What are you submitting, and where?
Bring the submission date and your current file. We will build the product version, place the documents under the 510(k) or Annex II sections, take the snapshot and download it.
Part of Regulated Product Lifecycle. See everything it does.
Less time on documentation. More time on the device.